Scientists from the Institute of Metal Research, Chinese Academy of Sciences (IMR, CAS), in collaboration with Peking University and Chongqing University, have clarified how the geometry and electronic properties of sub-nanometer Rh active sites govern C–H activation, product desorption and C–C bond cleavage in butane dehydrogenation. The work, published in ACS Catalysis, shows that fully exposed low-coordination Rh₃ clusters achieve a better balance between dehydrogenation activity and butene selectivity, offering a new strategy for designing efficient alkane dehydrogenation catalysts.
Butenes are key feedstocks for high-value chemicals, and butane dehydrogenation is an important route to produce them. However, alkane dehydrogenation is thermodynamically limited and typically requires high temperatures, which promote deep dehydrogenation, C–C bond cracking, carbon deposition and metal sintering. A central challenge is therefore to accelerate C–H activation while facilitating olefin desorption and suppressing cracking.
Rh exhibits strong C–H activation ability, but its catalytic performance is highly sensitive to active-site structure. Isolated Rh atoms can suffer from insufficient dehydrogenation activity, whereas larger Rh particles tend to promote C–C bond cleavage and side reactions. To resolve this contradiction, the team constructed Rh₁, Rh₂, Rh₃ and Rh₆ models on defective graphene, together with stepped Rh(211) and flat Rh(111) surfaces representing Rh nanoparticles, and combined density functional theory calculations with experiments.
The calculations revealed a volcano-type relationship between dehydrogenation activity and Rh size, with fully exposed Rh₂ and Rh₃ clusters showing more favorable thermodynamics. Rh atoms in these small clusters possess higher d-orbital energy, which favors transition-state binding during C–H activation while inhibiting C–C bond cleavage, so that butene tends to desorb rather than crack. On stepped Rh(211), the butene desorption energy is comparable to the C–C activation barrier, indicating facile C–C cleavage and a likely site for carbon deposition.
Guided by these predictions, the team prepared Rh single atoms, atomically dispersed fully exposed Rh₃ clusters, larger Rh clusters and nanoparticles on defect-rich graphene by tuning Rh loading. In butane dehydrogenation, the Rh₃ catalyst achieved an initial butane conversion of 10.3% and a C₄ dehydrogenation product selectivity of 89.5%. Kinetic and temperature-programmed experiments confirmed that Rh₃ clusters outperformed other Rh sizes, with activity following a volcano-type curve. Rh nanoparticles produced far more C₁–₃ products and suffered rapid deactivation and much higher carbon deposition, consistent with the theoretical predictions.
This work establishes the relationship between Rh active-site structure and catalytic performance, demonstrating that low-coordination, fully exposed Rh clusters can simultaneously promote dehydrogenation and suppress cracking and coking. It provides a new basis for active-site design in efficient alkane dehydrogenation catalysts.
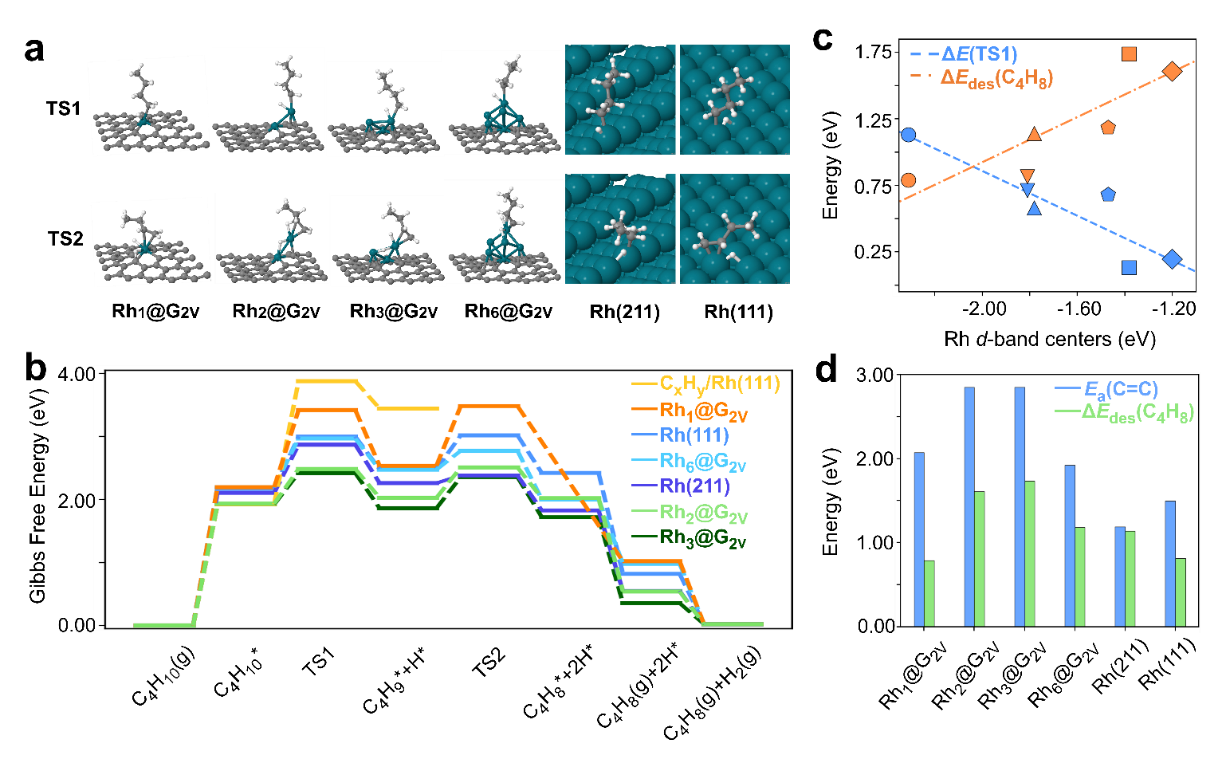
Theoretical calculation results. (a) Configurations of TS1 and TS2; (b) Gibbs free energy profiles; (c) Relationship among d-band center, butene adsorption energy and the first C–H activation barrier; (d) Comparison of butene adsorption energy and C=C activation energy for different configurations. (Image by IMR)
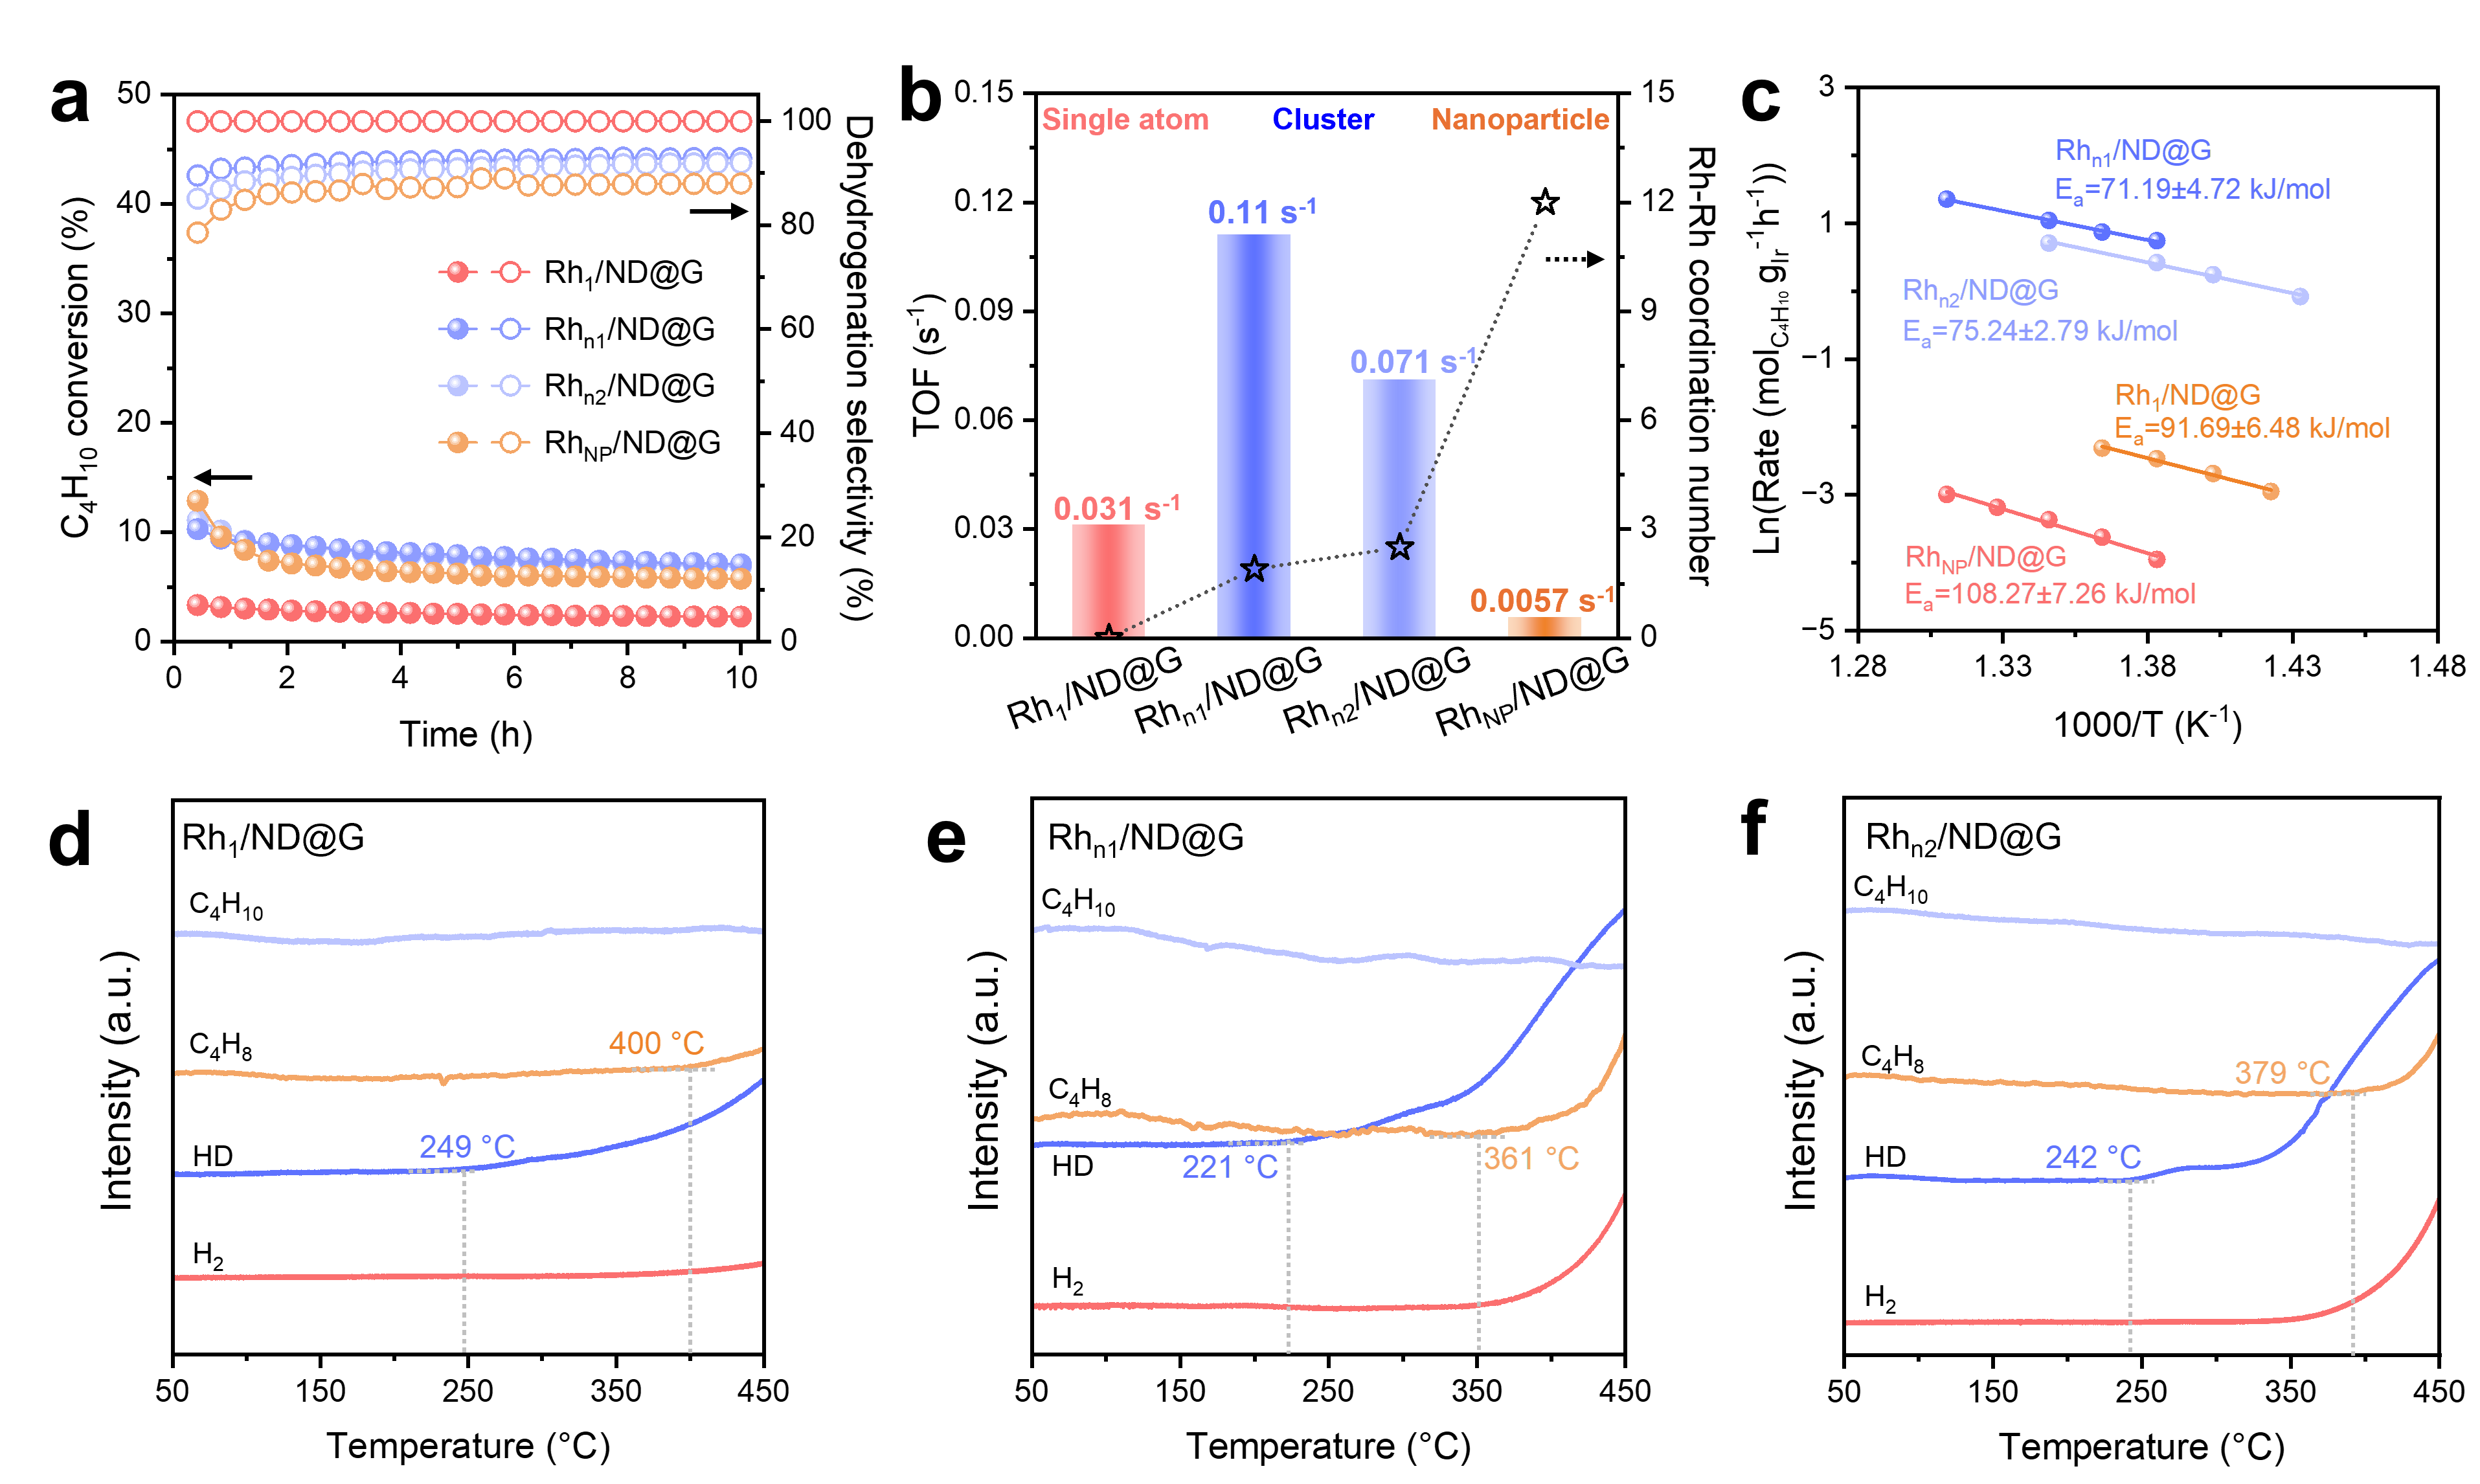
Butane dehydrogenation performance tests. (a) Butane conversion and C₄ olefin selectivity; (b) Relationship between TOF and the coordination number of Rh–Rh bonds; (c) Apparent activation energy measurements; (d)–(f) Temperature-programmed experimental results. (Image by IMR)
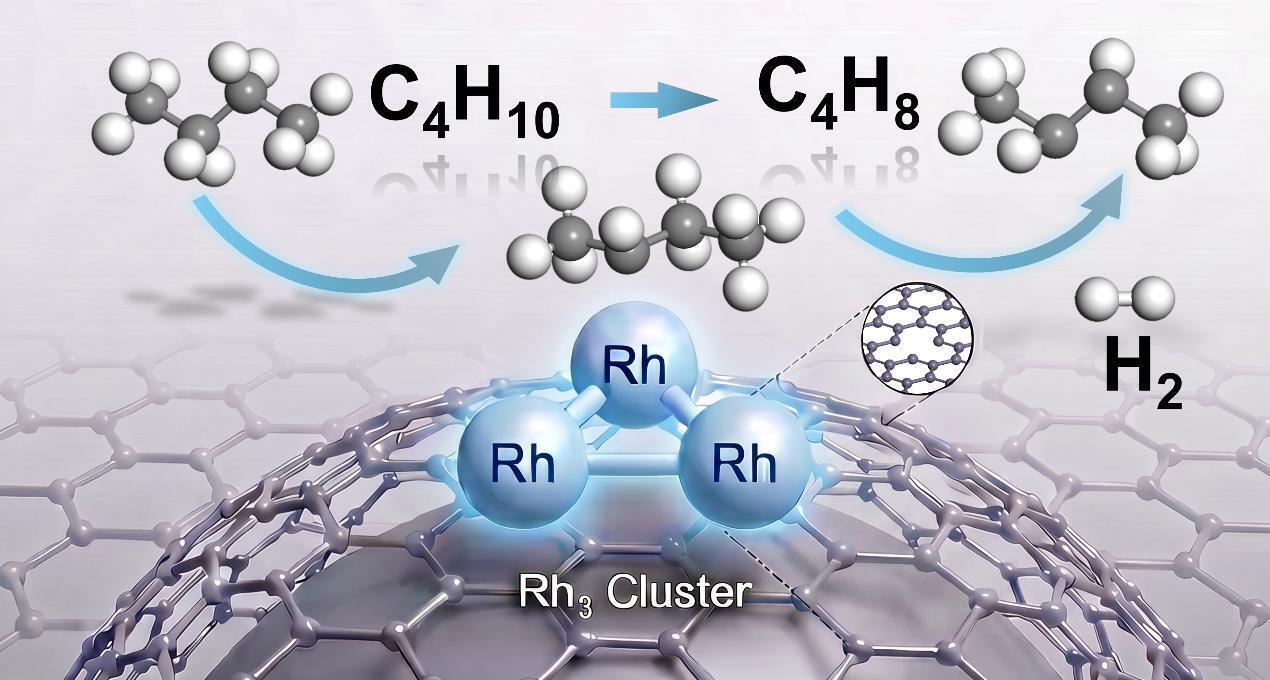
Schematic illustration of butane dehydrogenation over fully exposed Rh₃ clusters. (Image by IMR)